
Reviewed October 31, 2026 · For B2B procurement and development teams
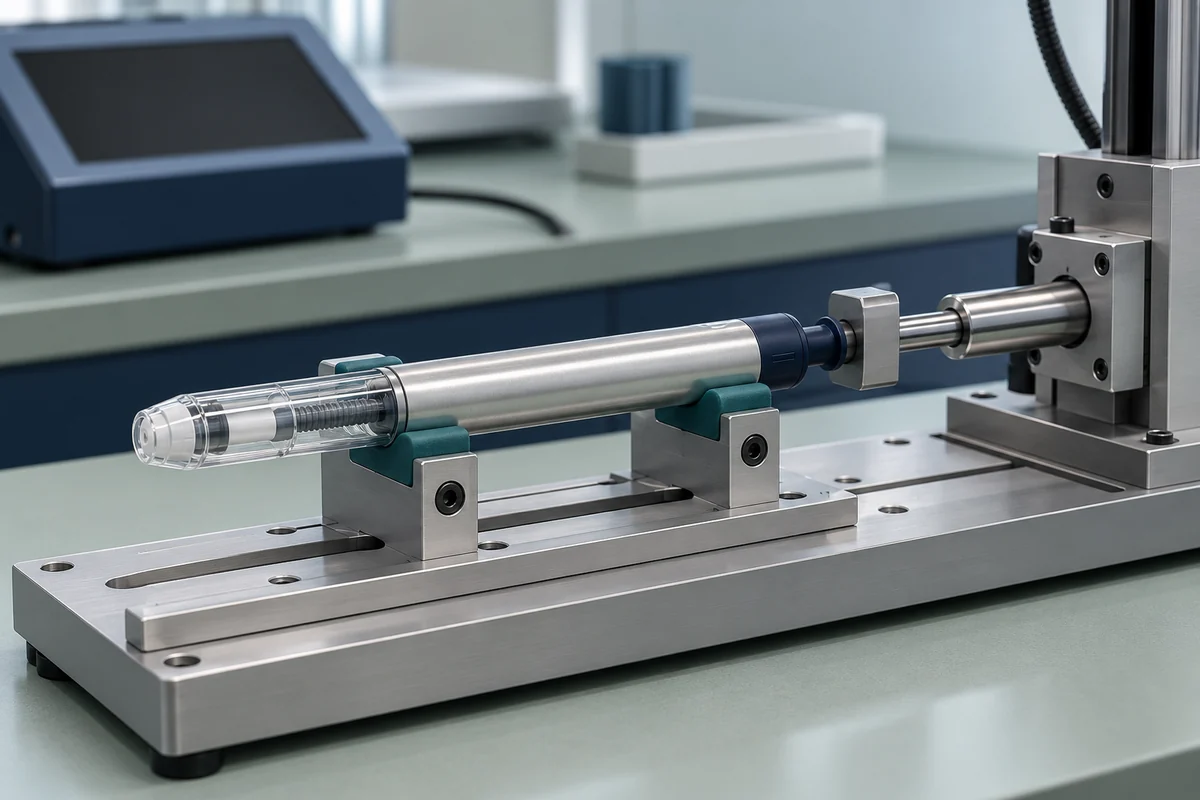
Injection pen activation force is the force a user applies to start or continue a defined device action, such as pressing a dose button or triggering an automated sequence. A useful result identifies the exact action, device state, test direction, loading rate, fixture alignment, environmental condition and lifecycle point. The peak value alone is rarely enough. A force-displacement trace can show preload, breakaway, travel, mechanism transitions and an abnormal interruption that one reported maximum would hide. ISO 11608-1:2022 provides requirements and test methods for needle-based injection systems, while FDA human-factors guidance places the user interface within a risk-based evaluation. Neither source publishes one universal activation-force limit for every pen. The project team should set acceptance criteria from the intended users, use conditions, device architecture and risk analysis before testing begins.
Scope boundary: This guide covers engineering evidence for a controlled injection-pen configuration. It does not set a clinical requirement, certify a product or replace the applicable standard, risk analysis or regulatory review.
Project inputs to control
- Defined user action and start/end points
- Device configuration, container and needle state
- Loading direction, rate and fixture alignment
- Temperature, conditioning and lifecycle point
- Force sensor range, resolution and calibration
- Trace processing, invalid-run and retest rules
- User population and critical-task rationale
- Acceptance criteria approved before testing
What does an activation-force result actually describe?
The result describes one controlled device action, not the effort required for every step in the injection sequence.
A manual dose button, an autoinjector trigger and a needle-shield release use different mechanisms and user motions. Name the action before assigning a requirement. Record where force is applied, the direction of travel and the event that marks activation. The same pen can produce different traces when it is primed, near end of content, fitted with another container or tested after mechanical conditioning.
Report the full trace where it helps explain the action. Preload, breakaway, a spring release and continuing travel can appear as separate features. If the analysis reduces that sequence to one number, the method should state which peak or interval it uses and why. Keep units, sign convention, zeroing and filtering rules with the result. That lets another reviewer distinguish mechanism behavior from fixture contact or data processing.
Terms to define in the protocol
- Force application point and direction
- Activation event and travel interval
- Peak, average or other reported trace feature
- Device state before and after the action
How should the fixture and test method be specified?
The fixture should reproduce the intended load path without adding side load, housing restraint or contact geometry that changes the mechanism response.
Describe the actuator tip, supports, alignment and loading rate. A hard narrow tip can contact a curved button differently from a finger pad, while excessive body clamping can stiffen the housing. Photograph the setup and retain fixture drawings or coordinates so a later test can reproduce it. The force sensor must have suitable range, resolution and calibration for the expected signal.
Predefine device conditioning, dwell time and test order. Temperature, storage, transport conditioning, container installation, needle attachment and mechanism cycling can affect the result. State whether the same unit provides several measurements or each unit is tested once. Repeated operation can change friction or consume a single-use mechanism. Separate setup checks from reportable runs, and write invalid-run and retest rules before data review. The method should preserve unusual traces rather than discard them because their peak falls inside a summary limit.
Method records buyers should request
- Fixture drawing and setup photograph
- Actuator speed and sampling rate
- Sensor identification and calibration status
- Conditioning, sequence and retest procedure
How should force-displacement data be reviewed?
Review the distribution of valid traces, the shape of each mechanism event and any failure mode alongside the predefined acceptance criteria.
A mean and standard deviation cannot show every relevant event. Plot individual traces or a justified overlay, then inspect outliers, secondary peaks, interrupted travel and units that never reach the defined activation point. Link each result to the device serial or sample identifier, component lots, configuration revision and conditioning state. If filtering or smoothing changes the reported peak, retain the raw data and document the processing method.
Statistical treatment should match the decision. Engineering characterization, design verification and lot release answer different questions. A small development study can expose mechanism behavior without supporting a population claim. A verification plan needs a sample rationale tied to the requirement and risk. Avoid choosing a pass limit from the observed sample after testing. Where several device variants share a mechanism, document why the selected worst-case configuration represents each claimed variant and which interfaces remain outside that argument.
Minimum review package
- Raw and processed traces
- Sample and component-lot traceability
- Deviation and failure-mode record
- Statistical rationale and scoped conclusion
How does bench force evidence connect to human factors?
Bench testing characterizes the device, while human-factors work evaluates whether intended users can complete the task safely in the intended use environment.
FDA's human-factors guidance asks manufacturers to identify user-interface risks and evaluate critical tasks. A force requirement should therefore trace to the user action and possible harm if the action is missed, delayed or performed incorrectly. Bench equipment cannot represent grip, posture, vision, dexterity, instructions or stress. Human-factors results also do not replace a controlled force measurement because they do not define the mechanism's engineering distribution.
Use both evidence streams to review the design. If users change grip, press the wrong surface or stop before activation, the team may need to revise geometry, feedback, labeling or the test requirement. Record which intended user groups and conditions support the requirement. For an OEM request, provide the action, user population, target use environment, device state and existing evidence. The supplier can then propose a method without assuming that a force value from another platform applies to the new program.
Questions for the design review
- Which user task does the requirement support?
- What happens if activation is incomplete?
- Which feedback confirms successful activation?
- What design change would trigger re-evaluation?
Evidence review matrix
| Evidence element | What to record | Buyer review question |
|---|---|---|
| Action definition | Load point, direction, start and completion event | Is the result tied to one observable device action? |
| Test setup | Fixture, actuator tip, rate, sensor and alignment | Could the setup add or remove force? |
| Sample state | Configuration, lots, conditioning and lifecycle | Which proposed units does the evidence represent? |
| Data package | Raw traces, processing, deviations and statistics | Can unusual behavior be reviewed? |
| Human-factors link | Critical task, users and use environment | Does the engineering requirement support the real task? |
Sources and related engineering guides
Use the current official source, its revision status and the controlled project configuration when converting this guide into requirements. Public summaries identify scope; they do not replace the full standard, qualified regulatory review or product-specific evidence.
Primary sources
- ISO 11608-1:2022, Needle-based injection systems, Part 1
- FDA, Applying Human Factors and Usability Engineering to Medical Devices
- FDA, Application of Human Factors Engineering Principles for Combination Products
Continue the topic cluster
- Injection pen human factors and usability
- Injection pen dose accuracy testing
- Injection pen design changes
- OEM/ODM injection pen engineering
Questions buyers also ask
No. The responsible team must define a product-specific requirement from the intended users, task, architecture and risk analysis, then apply the relevant standard and regulatory framework.
No. Activation force describes user or mechanism input for a defined action. Dose accuracy describes the delivered output. The two can interact but require separate requirements and evidence.
An average may be part of the summary, but it can hide failed activations, unusual trace shapes and variant behavior. Request individual traceability and the predefined acceptance method.
Include lifecycle and conditioning states that the risk analysis and intended claim require. The protocol should justify the selected states rather than apply one generic sequence to every device.