
Reviewed October 23, 2026 · For B2B procurement and development teams
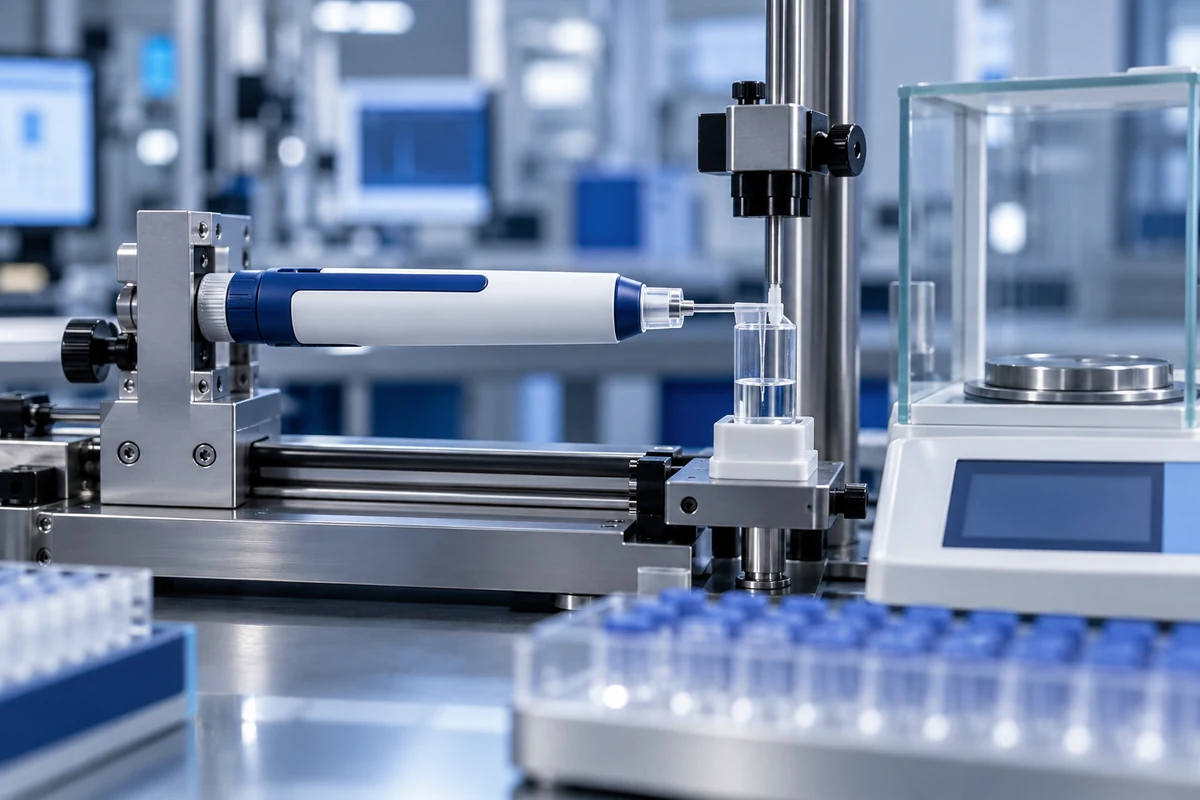
Injection pen dose accuracy cannot be established by one bench result or a nominal mechanism calculation. A defensible verification plan begins with the intended dose range, increments, primary container, fluid properties, needle configuration, environmental limits and lifecycle state. It then defines representative samples, preconditioning, measurement uncertainty, delivery procedure, statistical treatment and acceptance criteria before testing starts. ISO 11608-1:2022 is the central international reference for requirements and test methods for needle-based injection systems, while FDA’s pen-injector guidance explains the performance information expected for drug and biological product delivery systems. The exact protocol still belongs to the responsible manufacturer and product sponsor because the standard does not replace product-specific risk analysis, combination-product strategy or market submission requirements. HYTANBIO can support device fixtures, engineering samples and verification planning; final acceptance and regulatory conclusions must be tied to the controlled drug-device configuration.
Scope boundary: This page is a B2B engineering and evidence-planning guide. It is not medical advice, regulatory approval, a drug offer or a compatibility statement for an unnamed configuration.
Decision inputs to control
- Dose range, increments and end-of-content state
- Controlled primary container and stopper tolerances
- Test fluid density, viscosity and temperature
- Needle, priming, orientation and hold-time instructions
- Lifecycle, transport and environmental conditioning
- Measurement uncertainty and equipment suitability
- Sample-size and statistical rationale
- Deviation, retest and change-control rules
Define the measurand before selecting a test method
Dose accuracy is a system output, so the protocol must state exactly what is being measured and under which conditions. Inputs normally include the minimum, intermediate and maximum selectable dose; fixed-dose volume where relevant; dose increment; container fill state; fluid density or viscosity; needle specification; priming instruction; hold time; injection orientation; temperature and humidity; and whether the unit is new, conditioned or at a defined lifecycle point. Gravimetric measurement may be converted to volume only when the fluid density and balance performance are controlled. The test method should address evaporation, retained droplets, incomplete delivery and operator timing. For multi-dose pens, first, middle and last usable doses may create different force and travel conditions. For single-dose systems, activation, needle insertion, delivery completion and shield deployment can affect the result. These inputs should be traced to design requirements and risk controls before sample size or acceptance limits are approved.
Build a statistically defensible verification matrix
A verification matrix should connect each design requirement to a test condition, sample rationale, method, acceptance criterion and record owner. It should cover dimensional and material variants, container and stopper tolerance, fluid-property range, storage and transport conditioning, orientation, needle lots, mechanism lifecycle and foreseeable user-related variation that can affect delivery. The sampling plan must be justified for the claim being made; an engineering screening run is not the same as design verification. Measurement-system suitability includes calibration status, resolution, repeatability, reproducibility and uncertainty relative to the allowed error. Predefined rules are needed for invalid runs, device malfunctions, outliers and retests so results are not selected after the fact. When a platform is customized, the team should identify which legacy evidence remains applicable and which interfaces require new testing. The final report should preserve raw data, configuration identifiers, deviations and a conclusion limited to the tested design state.
Connect dose results to risk, usability and change control
A passing numerical result does not close every delivery risk. The team should review dose accuracy with activation force, dial or setting errors, priming, incomplete-dose feedback, needle attachment, hold time and user comprehension. The risk file should identify how an under-delivery, over-delivery, missed dose or unintended repeat dose could occur and which design or labeling controls address the sequence. Human-factors findings may trigger additional simulated-use conditions even when the basic bench method passes. Change control must define when new container lots, stopper coatings, lubricant, needle specification, spring, drive components, molded dimensions, assembly equipment or software require impact assessment and possible re-verification. FDA’s draft Essential Drug Delivery Outputs guidance can be used as a technical reference for identifying delivery-critical outputs, but it is explicitly a draft and not for implementation. Product sponsors should align the final evidence set with the current market pathway and submission strategy.
Related engineering guides
Cartridge Compatibility Guide · Human Factors Guide · OEM/ODM Engineering
Primary sources and status notes
Use the current official source and its revision status when converting this guide into project requirements. Standards summaries do not replace the purchased standard, and a source identified as draft is not binding or for implementation.
- ISO 11608-1:2022 — Needle-based injection systems, Part 1
- FDA — Technical Considerations for Pen, Jet, and Related Injectors
- FDA — Essential Drug Delivery Outputs, Draft Guidance, not for implementation
Questions buyers also ask
No. The protocol should cover the dose range and the lifecycle, container, environmental and use conditions that can affect delivery.
No. It provides requirements and test methods; conformity conclusions require controlled product-specific evidence and responsible-party review.
Not by itself. A surrogate can support engineering work only when its relevance to the final fluid and test objective is justified.
It should retain the tested configuration, method, equipment, raw data, acceptance criteria, deviations, statistics and a conclusion limited to that design state.