
Reviewed November 6, 2026 · For B2B procurement and development teams
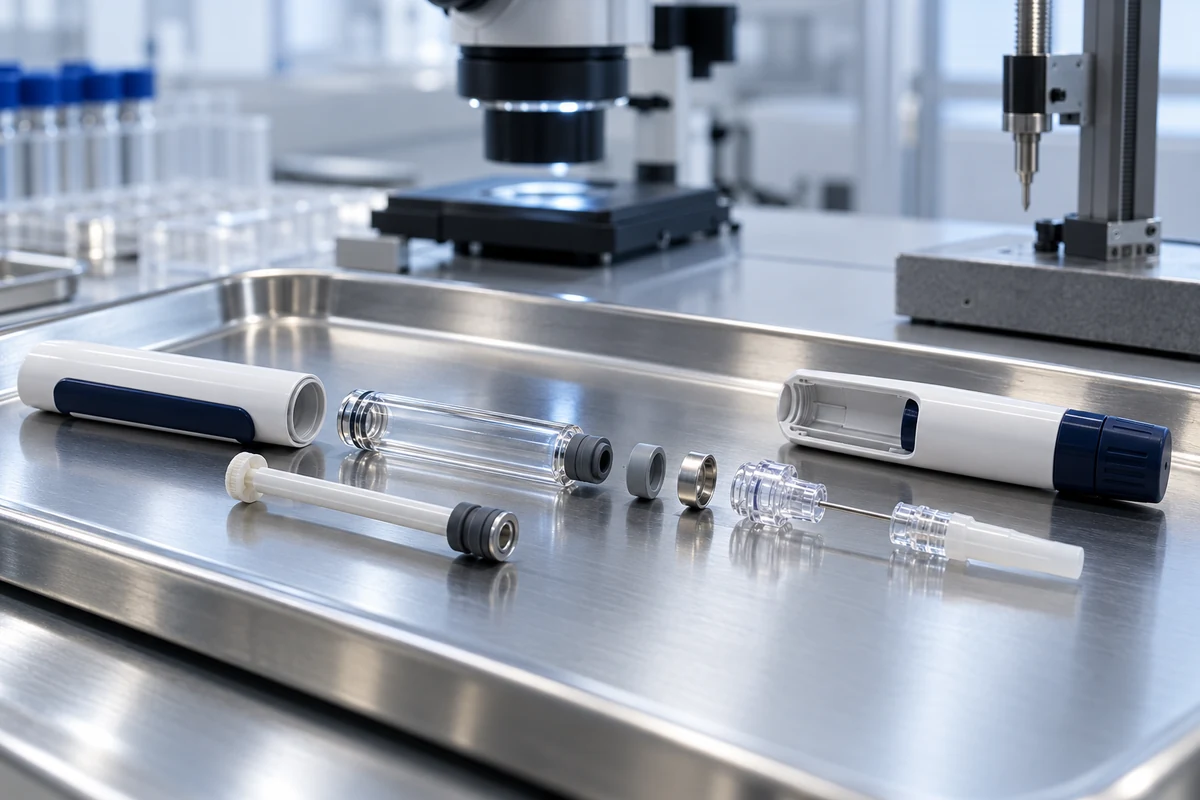
The primary container is a functional part of an injection pen system, not a generic package selected only by nominal volume. Cartridge barrel geometry, flange and neck dimensions, stopper design, lubrication, closure, septum, needle penetration, usable volume and component tolerances can change retention, drive force, leakage, dose accuracy and end-of-content behavior. ISO 11608-3:2022 addresses design-verification requirements and tests for containers and integrated fluid paths used with needle-based injection systems. A robust program therefore freezes the controlled container drawing and supplier configuration before mechanism tooling is finalized, then verifies the device and fluid path across representative tolerance, lifecycle, storage and transport conditions. HYTANBIO can evaluate mechanical interfaces and develop fixtures or samples, but the medicine sponsor remains responsible for formulation compatibility, container-closure integrity, stability, extractables and leachables, and the final drug-device evidence package.
Scope boundary: This page is a B2B engineering and evidence-planning guide. It is not medical advice, regulatory approval, a drug offer or a compatibility statement for an unnamed configuration.
Decision inputs to control
- Controlled cartridge or syringe drawing and revision
- Stopper, closure, septum and lubrication specification
- Dimensional tolerance and supplier-lot strategy
- Retention, alignment and plunger engagement
- Needle penetration and integrated fluid-path boundary
- Break-loose, glide and delivery-force range
- Leakage, dead volume and end-of-content behavior
- Pharmaceutical evidence and change-control ownership
Translate every container feature into an interface requirement
A nominal “3 mL cartridge” description is insufficient for design control. The interface specification should identify the barrel material and dimensions, neck and flange geometry, stopper material and profile, plunger contact surface, lubrication, crimp or cap construction, septum thickness, needle penetration zone, headspace, fill and usable volume, dimensional tolerances and supplier change-notification rules. Device requirements then define axial and radial location, retention, visibility, plunger engagement, available travel, anti-rotation features where relevant and protection from breakage or unintended movement. For reusable pens, the cartridge installation and removal sequence also becomes a user interface. Each dimension should have a source and a tolerance owner. If a public standard or supplier drawing leaves an interface open, the gap must be resolved by controlled engineering evidence rather than a nominal-size assumption.
Verify the container, mechanism and fluid path as one system
System verification should address the complete path from stored product to the delivery point. Depending on the architecture, this can include container retention, stopper movement, break-loose and glide force, septum piercing, needle connection, dead volume, leakage, air entry, blockage, particle generation, dose delivery, end-of-content behavior and post-use containment. Testing should represent container and device tolerance, relevant component lots, orientation, temperature, transport conditioning, storage time and lifecycle state. ISO 11608-3 provides the container and integrated-fluid-path framework, while ISO 11608-1 addresses the broader needle-based injection system. The project protocol still needs configuration-specific acceptance criteria and a rationale for any surrogate fluid or unfilled article. A mechanical fit check or successful assembly trial does not establish container-closure integrity, pharmaceutical compatibility, sterility maintenance or shelf life.
Separate device evidence from pharmaceutical evidence
The evidence plan should show where device engineering ends and pharmaceutical responsibility begins. A device supplier can measure dimensions, retention, drive force, leakage under defined methods, mechanical robustness and delivery outputs for controlled samples. The medicine sponsor typically owns formulation stability, container-closure suitability, material compatibility, extractables and leachables, sterility strategy, shipping validation and the final market submission. These workstreams interact: a stopper or lubricant change can alter both pharmaceutical quality and device force, while a needle or septum change can affect penetration, leakage and delivery. Quality agreements should define sample custody, test-article release, deviations, data access and change notification. Before design freeze, the team should issue an interface-control document and a responsibility matrix linking every critical container and fluid-path characteristic to its evidence owner and approval record.
Related engineering guides
Cartridge Compatibility Guide · Pen Needle Compatibility · Reusable Injection Pens
Primary sources and status notes
Use the current official source and its revision status when converting this guide into project requirements. Standards summaries do not replace the purchased standard, and a source identified as draft is not binding or for implementation.
- ISO 11608-3:2022 — Containers and integrated fluid paths
- FDA — Technical Considerations for Pen, Jet, and Related Injectors
- FDA — Container Closure Systems for Packaging Human Drugs and Biologics
Chemical evidence for the fluid-path configuration
Mechanical fit and delivery testing do not establish material compatibility by themselves. The companion guide, Injection Pen Extractables and Leachables: Material Risk Assessment, covers contact mapping, extractables study design, leachables assessment, toxicological ownership and supplier-change triggers for the finished drug delivery configuration.
Questions buyers also ask
No. Critical geometry, stopper, closure, lubrication and tolerances can differ and must be assessed against the exact pen design.
No. Mechanical fit does not establish material compatibility, stability, sterility or container-closure integrity.
It is the set of components through which the product travels as part of the delivery system, assessed with its actual interfaces and use sequence.
Assess re-verification after changes to the container, stopper, closure, lubricant, needle, device geometry, assembly process or relevant supplier.