
Reviewed October 23, 2026 · For B2B procurement and development teams
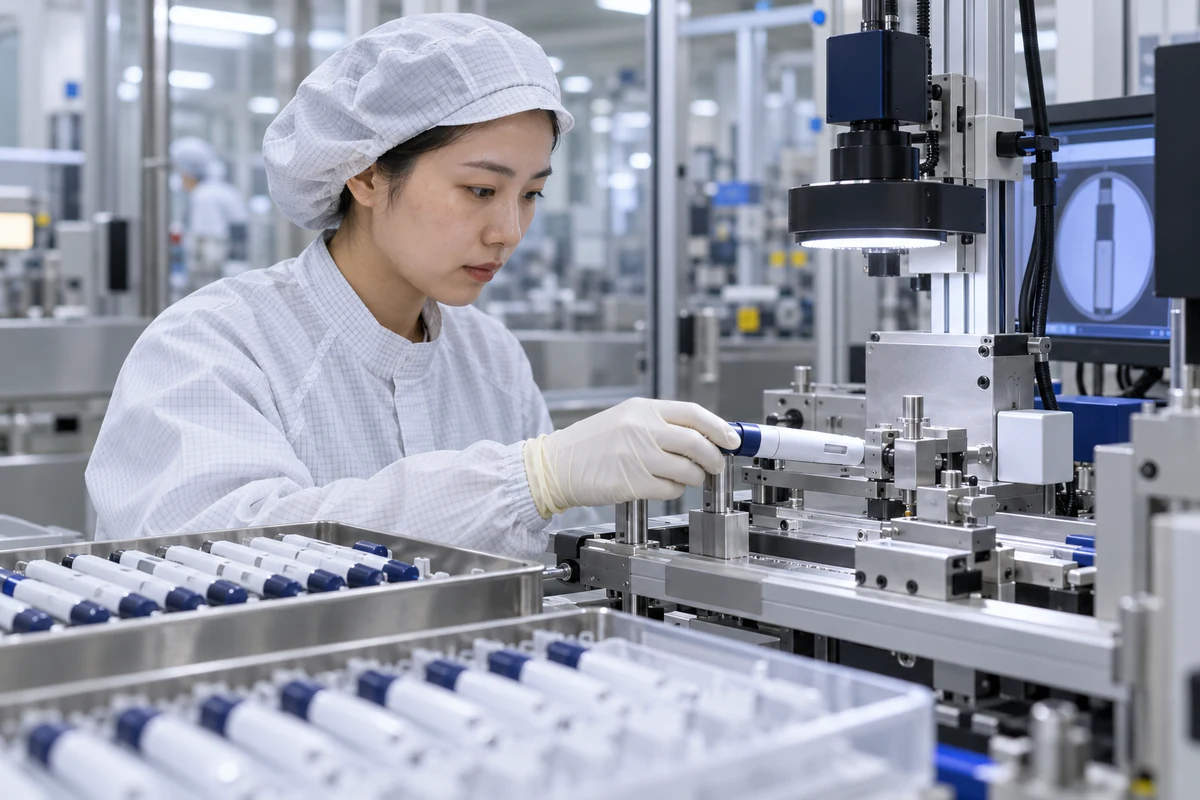
Injection pen design transfer is complete only when approved design outputs can be reproduced through controlled production specifications, equipment, inspection methods, work instructions and supplier controls. A pilot build that produces acceptable samples is useful evidence, but it does not by itself demonstrate a capable and validated manufacturing process. FDA’s Quality Management System Regulation became effective on February 2, 2026 and incorporates ISO 13485:2016 by reference into 21 CFR Part 820. For an injection pen program, the transfer plan should connect design inputs, risk controls and verification results to the bill of materials, drawings, tooling, assembly parameters, software or firmware where applicable, acceptance activities, traceability and change control. Processes whose results cannot be fully verified by subsequent inspection require an appropriate validation strategy. HYTANBIO can support industrialization and manufacturing evidence; the responsible manufacturer must approve the quality-system scope and finished-product release strategy.
Scope boundary: This page is a B2B engineering and evidence-planning guide. It is not medical advice, regulatory approval, a drug offer or a compatibility statement for an unnamed configuration.
Decision inputs to control
- Released bill of materials, drawings and specifications
- Risk-control to production-control traceability
- Tooling, fixtures, software and equipment configuration
- Measurement-system and inspection-method suitability
- Process-validation rationale and protocol
- Supplier qualification and incoming controls
- Training, traceability and release records
- Deviation, change and revalidation triggers
Create a design-transfer package that production can execute
The transfer package should define more than final drawings. It normally includes the approved bill of materials, material and component specifications, tolerances, tooling and fixture drawings, assembly sequence, parameter windows, inspection methods, calibration needs, software versions where relevant, packaging specifications, labeling inputs, traceability rules and training requirements. Each item should link back to a design output, risk control or acceptance requirement. Production-equivalent units are needed to confirm that specifications are understandable and achievable outside the development bench. The team should review supplier capability, component qualification, measurement-system suitability and incoming controls before routine manufacture. Open deviations, temporary tooling or manual adjustments must be visible rather than absorbed into an informal process. A formal transfer review should identify the released revision, responsible site, unresolved risks, approval records and conditions that must be met before process qualification or commercial release.
Validate processes where inspection alone cannot assure the result
Process validation should be driven by process risk and the ability of later inspection to verify the output. Injection pen examples may include molding characteristics, bonding, welding, lubrication, spring or drive assembly, automated dosing-mechanism assembly, software-controlled stations, sealing and packaging operations. The validation strategy should define equipment qualification, operating ranges, challenge conditions, material and component lots, sampling, acceptance criteria, data handling, deviations and revalidation triggers. Critical parameters should be linked to critical quality or performance characteristics rather than chosen only because the machine records them. A successful design-verification report does not validate the manufacturing process, and final inspection cannot always detect latent bond, weld, lubrication or assembly defects. The quality plan should also define preventive maintenance, calibration, line clearance, operator qualification, in-process controls and statistical monitoring needed to keep the validated state.
Use QMSR, ISO 13485 and ISO 14971 as a connected system
FDA’s current QMSR aligns Part 820 more closely with ISO 13485:2016, while FDA inspection and U.S.-specific statutory requirements still apply. ISO 14971:2019 provides the lifecycle risk-management framework and was confirmed as current by ISO in 2025. For a transferred injection pen, risk controls must remain connected to supplier controls, process parameters, inspection, nonconformance handling, complaint information and production or post-production feedback. The manufacturer should define how changes to components, molds, software, equipment, site, test methods or process windows are assessed before implementation. FDA’s QMSR frequently asked questions note that the inspection approach changed with the regulation and that records previously exempt from review under former provisions may now be inspected. Teams should update procedures and training to the current framework rather than reusing an old Quality System Regulation checklist without review.
Related engineering guides
Manufacturing Process Guide · Factory and Quality Approach · OEM/ODM Engineering
Primary sources and status notes
Use the current official source and its revision status when converting this guide into project requirements. Standards summaries do not replace the purchased standard, and a source identified as draft is not binding or for implementation.
- FDA — Quality Management System Regulation
- FDA — QMSR Frequently Asked Questions
- ISO 13485:2016 — Medical devices quality management systems
- ISO 14971:2019 — Application of risk management to medical devices
Questions buyers also ask
FDA states that the Quality Management System Regulation became effective on February 2, 2026.
No. A pilot can support transfer, but validation needs predefined methods, acceptance criteria and evidence for the controlled production process.
Not when later inspection cannot fully verify the process result or detect latent defects.
Assess component, supplier, tool, process, software, site, test-method and specification changes before implementation.